Benchmark Proteomics Differential Expression Results
Source:R/Benchmarking_Single.R
benchmarking_proteomics.RdEvaluates the reliability of DE results by comparing them against known ground truth (multi-species spike-in experiments). Computes classification metrics, dispersion statistics, and generates visualizations.
Usage
benchmarking_proteomics(
  de_res,
  expected_values,
  alpha = 0.05,
  lfc_thr = 0,
  p_col = "adj.P.Val",
  comparisons = NULL,
  assay = NULL,
  output_dir = NULL,
  verbose = TRUE,
  species_colors = NULL,
  species_df = NULL
)
Arguments
- de_res
Data frame with DE results. Required columns: Protein.IDs, logFC, P.Value, adj.P.Val, Change, Comparison. Optional: Gene.Names, Assay, Species (if not using species_df).
- expected_values
Data frame with expected values. Required columns: Comparison, Species, expected_logFC.
- alpha
Significance threshold (default: 0.05)
- lfc_thr
Log fold-change threshold (default: 0)
- p_col
P-value column to use (default: "adj.P.Val")
- comparisons
Comparisons to include (NULL = all)
- assay
Assay to filter (NULL = all)
- output_dir
Directory for exported files (NULL = no export)
- verbose
Print progress messages (default: TRUE)
- species_colors
Named vector of colors per species (optional)
- species_df
Data frame with Protein.IDs and Species columns (optional). If NULL, de_res must already contain a Species column.
Value
List with:
metrics_table: Classification metrics per comparison
confusion_by_species: Confusion matrix per Comparison x Species
confusion_overall: Confusion matrix aggregated per Comparison
dispersion_metrics: Dispersion stats per Comparison x Species
classified_df: Full classified data frame
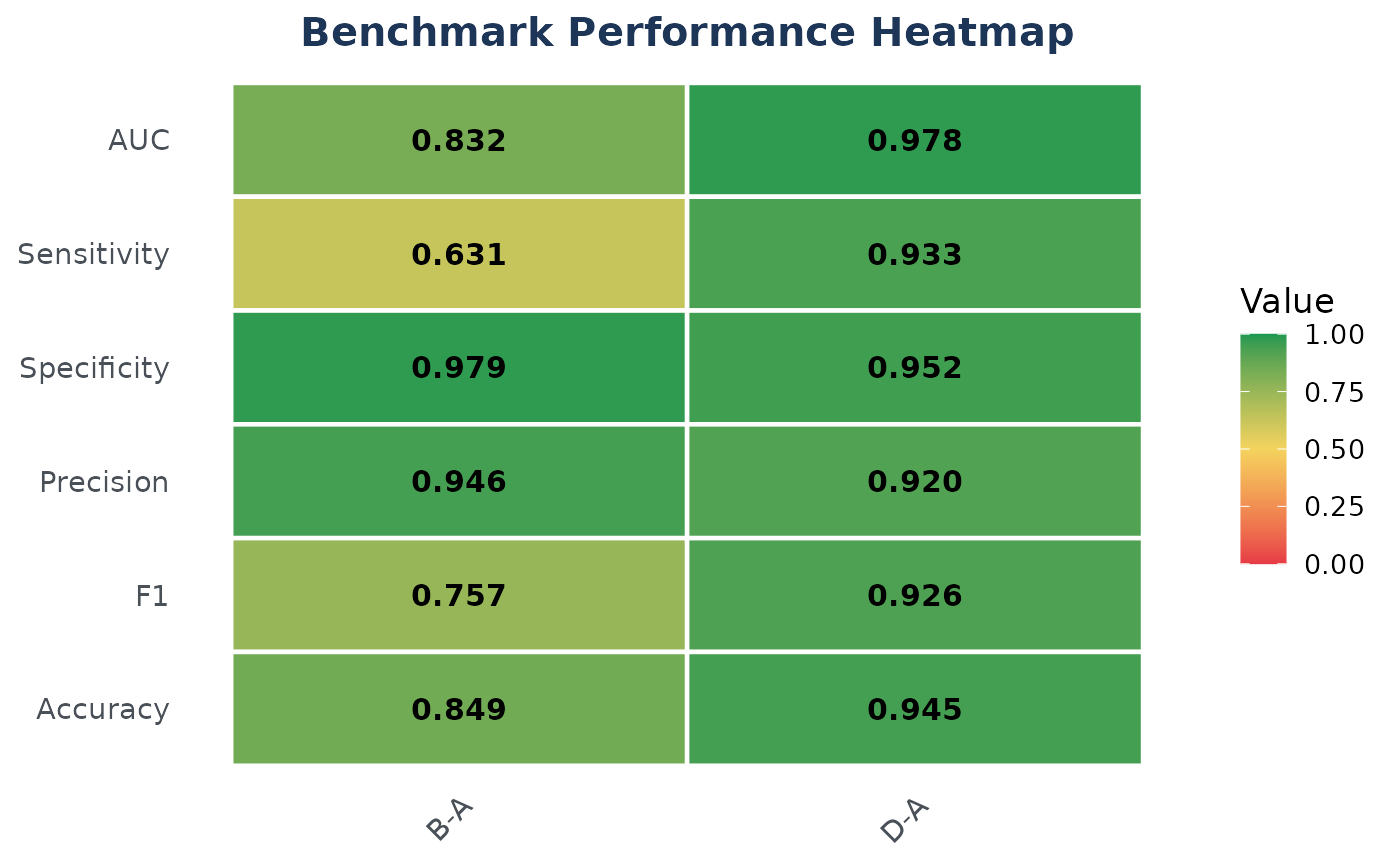
gg_heatmap: ggplot2 performance heatmap
gg_confusion_by_species: ggplot2 confusion matrix heatmap (by species)
gg_confusion_overall: ggplot2 confusion matrix heatmap (aggregated)
gg_auc_bars: ggplot2 AUC bar chart
gg_metrics_bars: ggplot2 grouped metrics bar chart
gg_signif_bars: ggplot2 significant proteins stacked bars (absolute)
hc_volcano_list: Named list of Highcharter volcano plots
parameters: List of parameters used
Examples
data(nadia_dia)
de <- process_proteomics(nadia_dia, verbose = FALSE)$DEPs_results
sp <- utils::read.delim(system.file("extdata", "nadia_dia_spikein.tsv.gz",
                                    package = "NADIA"))
species_df <- data.frame(Protein.IDs = sp$PG.ProteinGroups,
                         Species = sp$PG.OrganismId)
ev <- data.frame(Comparison = rep(c("B-A", "D-A"), each = 2),
                 Species = c("ECOLI", "YEAST"),
                 expected_logFC = c(1, -0.58, 2, -3.3))
bench <- benchmarking_proteomics(de, ev, species_df = species_df,
                                 verbose = FALSE)
bench$metrics_table
#>   Comparison  TP FP   TN  FN Sensitivity Specificity Precision    NPV Accuracy
#> 1        B-A 469 27 1227 274      0.6312      0.9785    0.9456 0.8175   0.8493
#> 2        D-A 693 60 1194  50      0.9327      0.9522    0.9203 0.9598   0.9449
#>       F1    MCC    AUC Performance
#> 1 0.7571 0.6821 0.8323        Good
#> 2 0.9265 0.8825 0.9779   Excellent
bench$gg_heatmap